# RhoFold+: Accurate RNA 3D structure prediction using a language model-based deep learning approach ![header](View.png) This is the open source code for RhoFold+.
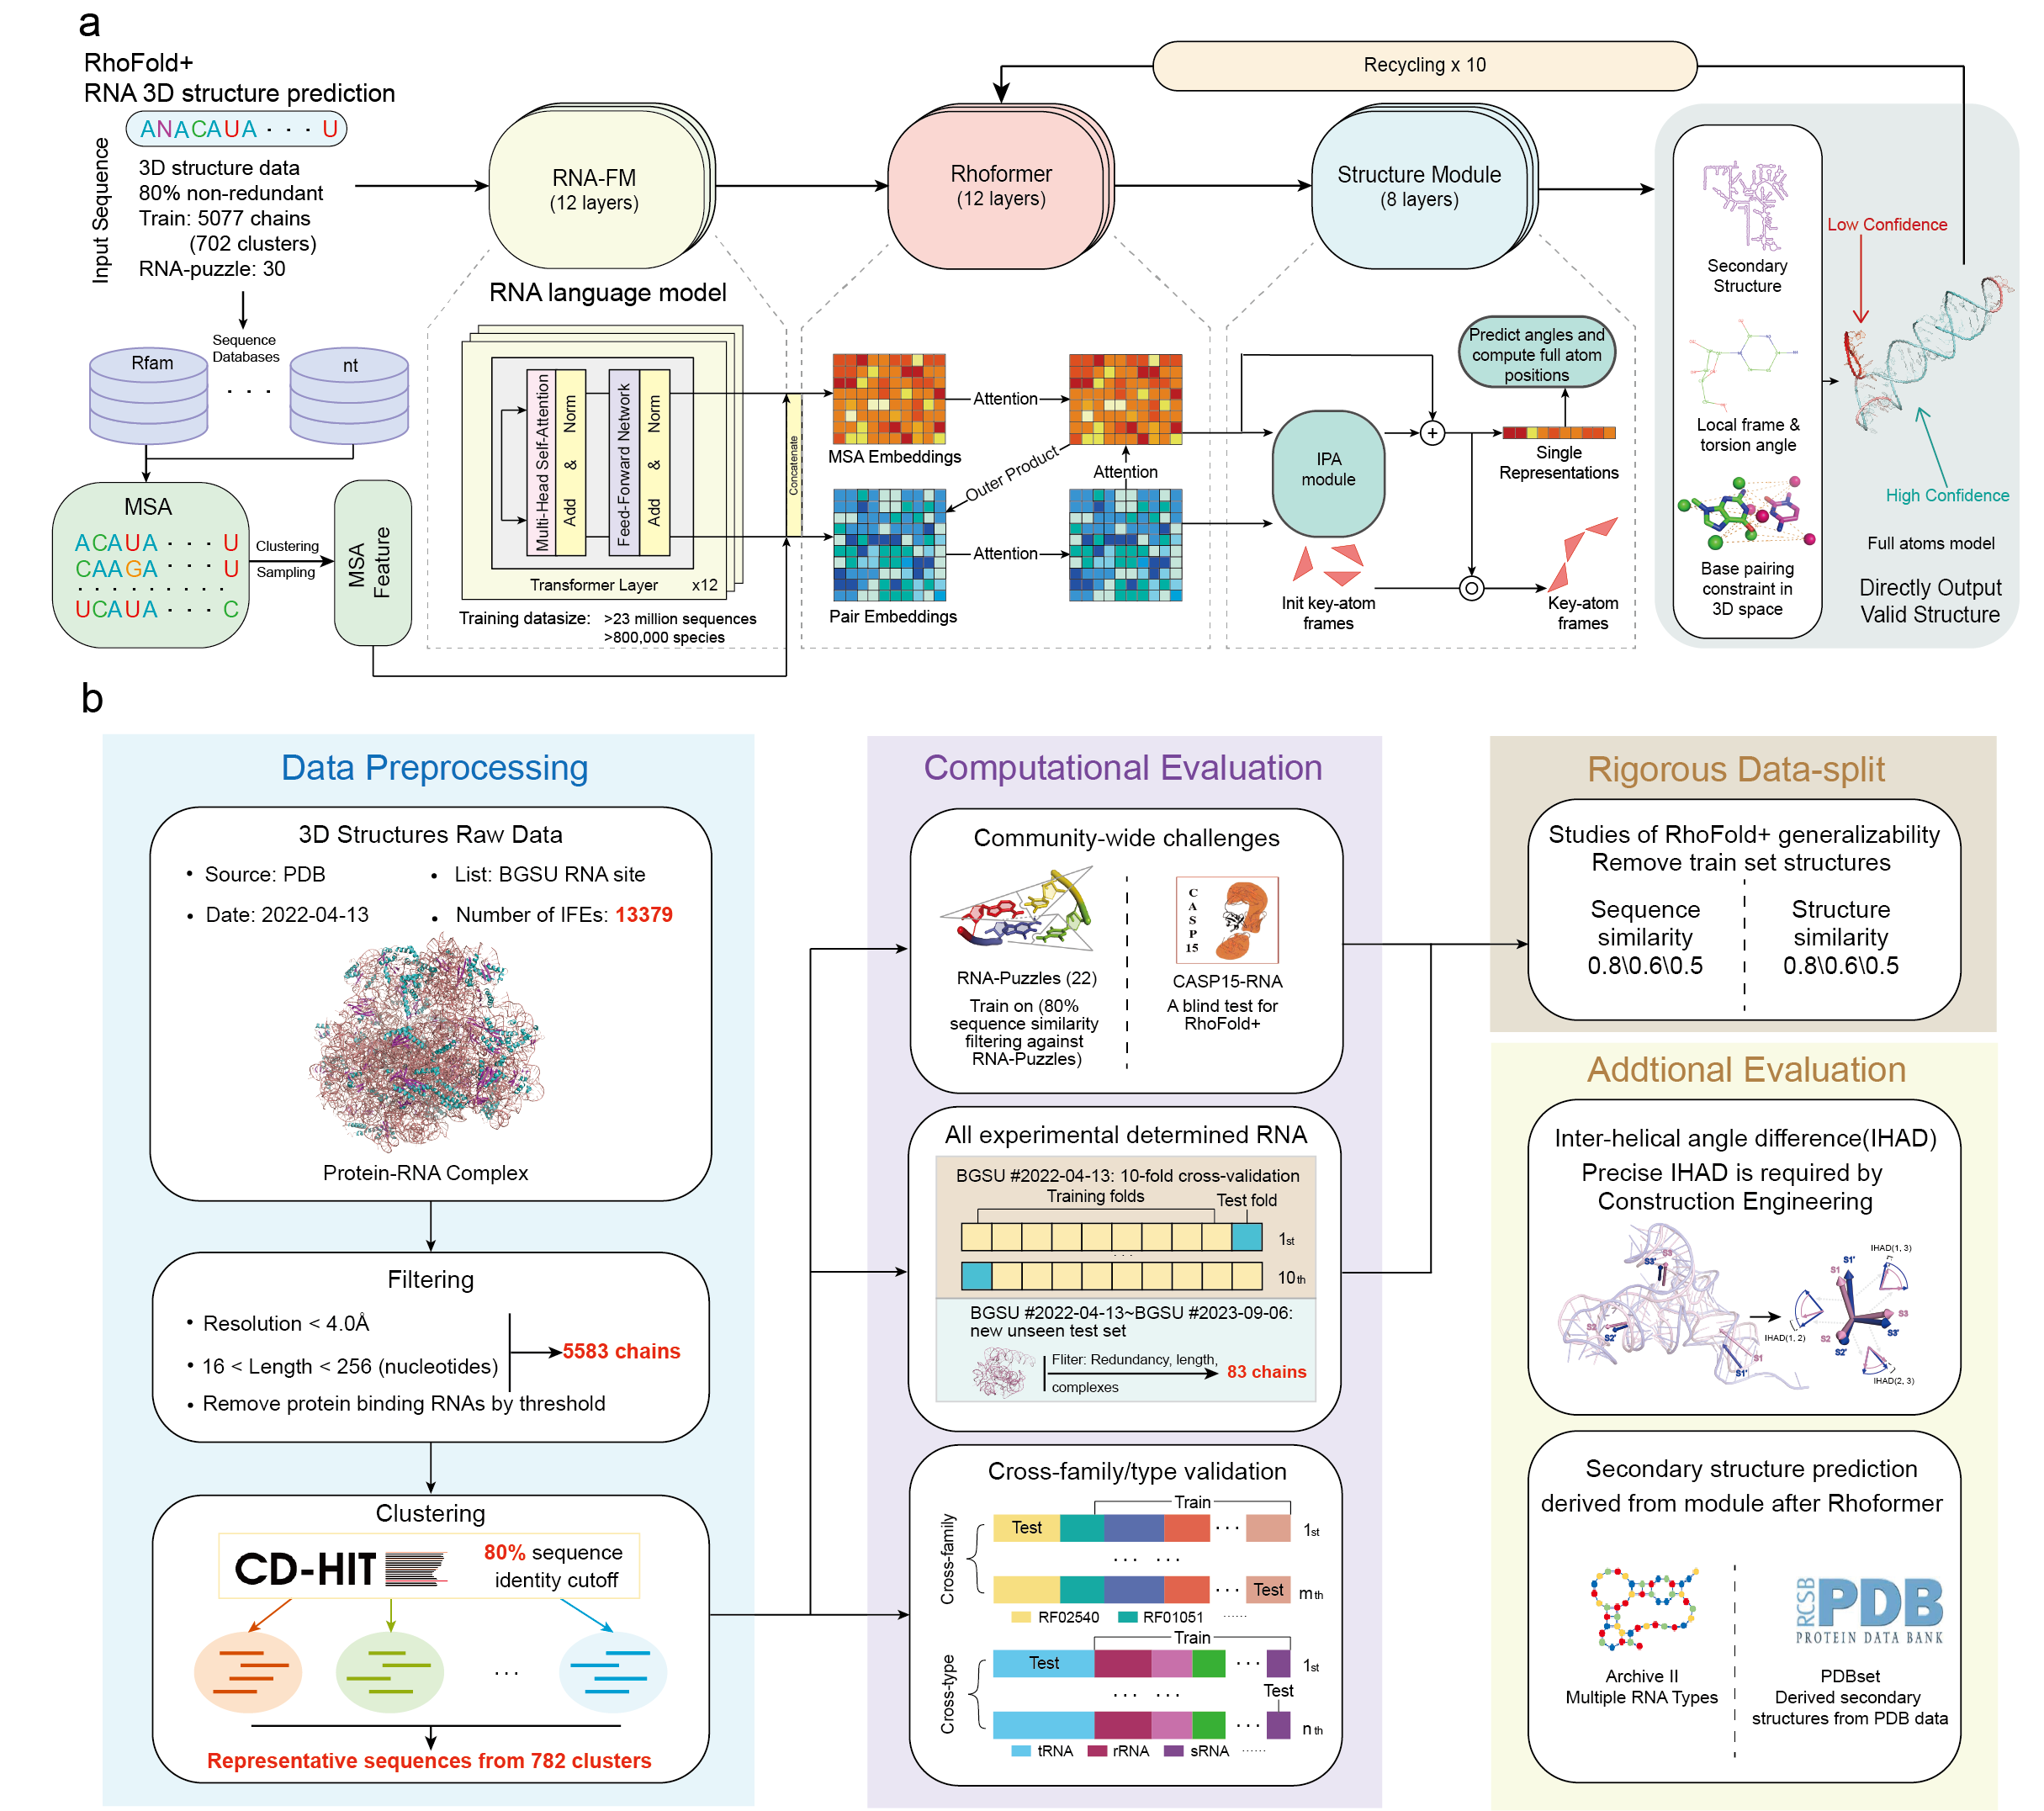
Citation ```bibtex @article{shen2022e2efold, title={E2Efold-3D: End-to-End Deep Learning Method for accurate de novo RNA 3D Structure Prediction}, author={Shen, Tao and Hu, Zhihang and Peng, Zhangzhi and Chen, Jiayang and Xiong, Peng and Hong, Liang and Zheng, Liangzhen and Wang, Yixuan and King, Irwin and Wang, Sheng and others}, journal={arXiv preprint arXiv:2207.01586}, year={2022} } ```
Table of contents - [Recent Updates](#New_Updates) - [Online Server](#Online_Server) - [Local Environment Setup](#Local_Environment_Setup) - [For Linux Users](#Installation_Linux) - [Download Pre-trained Model](#Pretrained_Model) - [Usage](#usage) - [Input Arguments](#Arguments) - [Output Files](#RhoFold_outputs) - [Examples](#Examples) - [Training Data](#Training_Data) - [Citations](#citations) - [License](#license)
## Updates *** Dec 31 / 2023 *** Integrated inferencing with clustered, sampled MSAs in RhoFold+. *** Oct 10 / 2023 *** Initial commits: 1. [Pretrained model](https://drive.google.com/file/d/1To2bjbhQLFx1k8hBOW5q1JFq6ut27XEv/view?usp=sharing) is provided. ## Online Server No need to create the environment locally, you can also access RhoFold+ easily through its online server: https://proj.cse.cuhk.edu.hk/aihlab/RhoFold/ ## Local Environment Setup For an updated step-by-step tutorial, we have also developed a protocol for RhoFold+ available at: https://github.com/WangJiuming/rhofold_protocol. To create environment with conda, first, download the repository and create the environment. ### Linux Users (MacOS is currently not supported) ``` git clone https://github.com/ml4bio/RhoFold.git cd ./RhoFold conda env create -f ./envs/environment_linux.yaml ``` Then, activate the "RhoFold" environment. ``` conda activate RhoFold python setup.py install ``` ### Download pre-trained model The model should be able to automatically download the pre-trained checkpoint before inference. Perform the following manual download in case the automatic download is unavailable. ``` cd ./pretrained wget https://huggingface.co/cuhkaih/rhofold/resolve/main/rhofold_pretrained_params.pt -O RhoFold_pretrained.pt cd ../ ``` ## Usage ### Input Arguments ```commandline python inference.py --input_fas INPUT_FAS Path to the input fasta file. Valid nucleic acids in RNA sequence: A, U, G, C. Input of sequence standalone is in testing. It's not as accurate as inputs of sequences combined with MSA. The former is only for the user to generate a quick reference structure. --input_a3m INPUT_A3M Path to the input msa file, default None. If --input_a3m is not given (set to None), MSA will be generated automatically. --output_dir OUTPUT_DIR Path to the output dir. Tertiary Structure prediction is saved in .pdb format (pLDDT score is recorded in the B-factor column). Distogram prediction is saved in .npz format. Secondary structure prediction is save in .ct format. --device DEVICE Default cpu. If GPUs are available, you can set --device cuda: for faster prediction. --ckpt CKPT Path to the pretrained model. Default ./pretrained/model_20221010_params.pt --relax_steps RELAX_STEPS Num of steps for structure refinement, default 1000. --single_seq_pred Default False. If --single_seq_pred is set to True, the modeling will run using single sequence only (input_fas) --database_dpath Path to the sequence database for MSA construction. Default ./database --binary_dpath Path to the executable. Default ./RhoFold/data/bin ``` ### Output Files The outputs will be saved in the directory provided via the `--output_dir` flag of `inference.py`. The outputs include the unrelaxed structures, relaxed structures, prediction metadata, and running log. The `--output_dir` directory will have the following structure: ``` <--output_dir>/ results.npz ss.ct unrelaxed_model.pdb relaxed_{relax_steps}_model.pdb log.txt ``` The contents of each output file are as follows: * `results.npz` – A `.npz` file containing the distogram prediction of RhoFold+ in NumPy arrays. * `ss.ct` – A .ct format text file containing the predicted secondary structure. * `unrelaxed_model.pdb` – A PDB format file containing the predicted structure from deep learning. * `relaxed_{relax_steps}_model.pdb` – A PDB format file containing the amber relaxed structure from unrelaxed_model.pdb. * `log.txt` – A txt file containing the running log. ### Examples Below are examples on how to use RhoFold+ in different scenarios. #### Folding with sequence and given MSA as input ```commandline python inference.py --input_fas ./example/input/3owzA/3owzA.fasta --input_a3m ./example/input/3owzA/3owzA.a3m --output_dir ./example/output/3owzA/ --ckpt ./pretrained/RhoFold_pretrained.pt ``` #### Folding with sampled, clustered MSA as input ```commandline python ./scripts/rhofold_msa_sampler_clust.py -i MSA_PATH -o OUT_DIR -n NUM_CLUST python inference.py --input_fas ./example/input/3owzA/3owzA.fasta --input_a3m OUT_DIR --output_dir ./example/output/3owzA/ --ckpt ./pretrained/RhoFold_pretrained.pt ``` #### Folding with single sequence as input 1.Sequence standalone This function is in testing. It's not as accurate as the MSA version. It's only for the user to generate a quick reference structure. ```commandline python inference.py --input_fas ./example/input/3owzA/3owzA.fasta --single_seq_pred True --output_dir ./example/output/3owzA/ --ckpt ./pretrained/RhoFold_pretrained.pt ``` 2.With our constructed MSA (**Full version of RhoFold+**) To support MSA construction, 3 sequence databases (RNAcentral, Rfam, and nt) totaling about 900GB need to be downloaded. **Warning**: You should ensure that there are adequate space for saving the data. Otherwise you can directly utilize our [online server](https://proj.cse.cuhk.edu.hk/aihlab/RhoFold/). ``` ./database/bin/builddb.sh ``` Then you can run the following command lines: ``` python inference.py --input_fas ./example/input/3owzA/3owzA.fasta --output_dir ./example/output/3owzA/ --ckpt ./pretrained/RhoFold_pretrained.pt ``` ## Training Data RhoFold's training data is available on the [Hugging Face repository](https://huggingface.co/datasets/cuhkaih/rhofold) for non-commercial use. To request access, users can submit an application through the provided form. [//]: # (AlphaFold's, OpenFold's and, by extension, xTrimoMultimer source code is licensed under the permissive Apache Licence, Version 2.0.) ## Citations ```bibtex @article{shen2024accurate, title={Accurate RNA 3D structure prediction using a language model-based deep learning approach}, author={Shen, Tao and Hu, Zhihang and Sun, Siqi and Liu, Di and Wong, Felix and Wang, Jiuming and Chen, Jiayang and Wang, Yixuan and Hong, Liang and Xiao, Jin and others}, journal={Nature Methods}, pages={1--12}, year={2024}, publisher={Nature Publishing Group US New York} } ``` ## License This source code is licensed under the Apache license found in the `LICENSE` file in the root directory of this source tree.