pymol-mcp
Headless PyMOL as an MCP server — drive molecular visualization, GROMACS/LAMMPS trajectories, and clathrate-hydrate cage science from your LLM.
Demo · Quick Start · Highlights · Features · Tools · Cage Science · 한국어


---
> [!NOTE]
> An MCP server that embeds **PyMOL in-process, headless** — no GUI, no socket plugin, no manual setup.
> It exposes 30+ **typed** tools, **returns rendered images inline** so the model can see what it draws,
> loads **GROMACS/LAMMPS** trajectories, and ships a **clathrate-hydrate analysis** toolkit
> (H-bond networks, F3/F4 order parameters) with **numerically validated** science.
## Demo
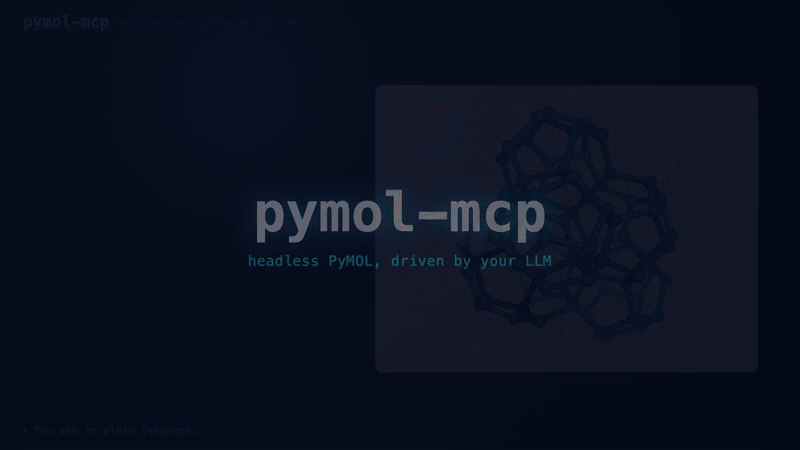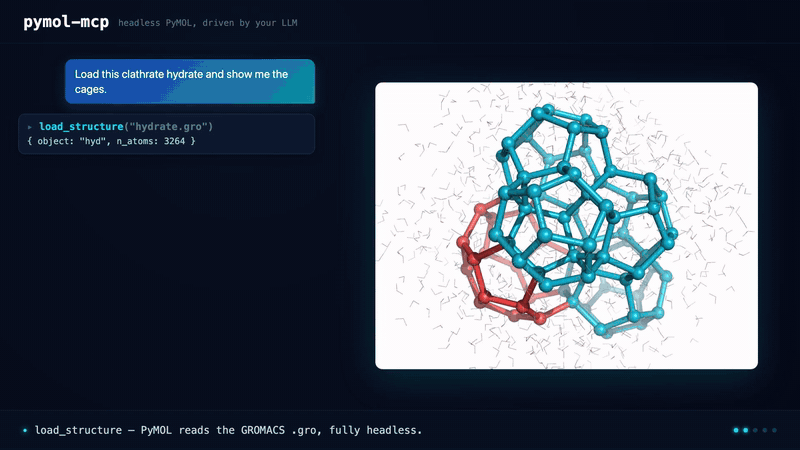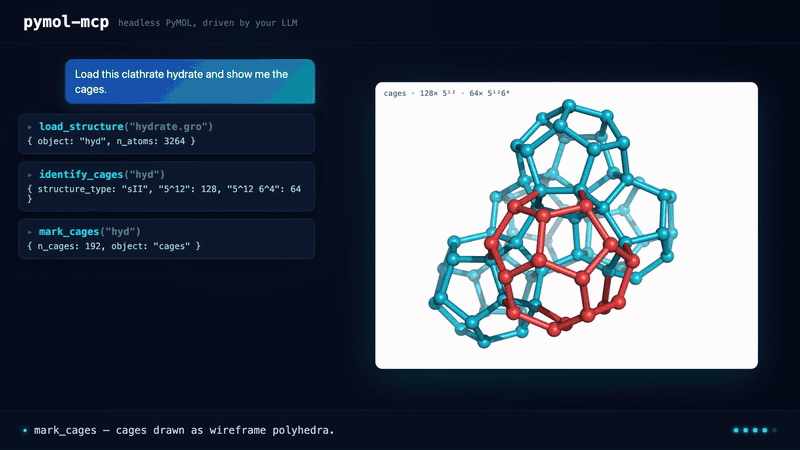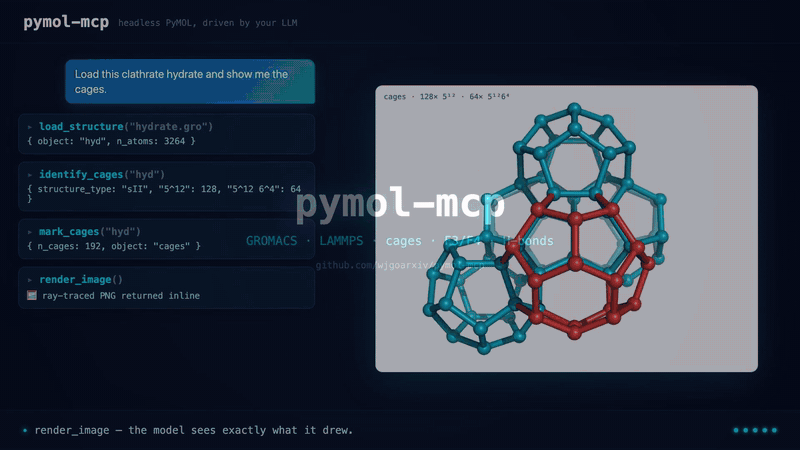
Ask in plain language → the model calls typed tools → headless PyMOL renders it. (full-quality MP4)
## Highlights
| | |
|---|---|
| **Runtime** | Embedded, **headless** `pymol2` — no GUI, no plugin, no socket |
| **Tools** | **30+ typed tools** with real, structured return values |
| **Vision** | ray-traced PNG **returned inline** so the model sees what it draws |
| **MD trajectories** | GROMACS `.xtc/.trr` + LAMMPS dump (MDAnalysis bridge) |
| **Domain science** | cage perception (TRACE), occupancy, H-bonds, F3/F4 — **validated** |
| **Robustness** | worker-thread session, **stdout-safe** transport, pytest suite |
| **Safety** | arbitrary-code passthrough **off by default** |
| **Directory** | [Glama](https://glama.ai/mcp/servers/wjgoarxiv/pymol-mcp) (`wjgoarxiv/pymol-mcp`, release v0.1.0) and [awesome-mcp-servers](https://github.com/punkpeye/awesome-mcp-servers) Biology ([PR #9110](https://github.com/punkpeye/awesome-mcp-servers/pull/9110), merged) |
## Features
- **Embedded & headless** — one long-lived PyMOL instance on a dedicated worker thread; nothing to click, works in CI.
- **The model can *see*** — `render_image` ray-traces and returns a PNG as MCP image content. Use `ray_style="pymolwiki_color"` for the PyMOLWiki quantized-color outline profile.
- **MD-native** — load GROMACS `.gro`+`.xtc`, or bridge LAMMPS/NetCDF/… through MDAnalysis with in-memory coordinate injection.
- **Clathrate-hydrate toolkit** — H-bond networks and **F3/F4** order parameters ported from a validated Rust engine, all in nm with correct triclinic PBC.
- **Typed, safe tools** — every argument is schema-validated; the arbitrary-code passthrough is **opt-in** (`PYMOL_MCP_ALLOW_CODE_EXEC=1`).
- **Protocol-hardened** — PyMOL's chatty stdout is permanently redirected so it can never corrupt the JSON-RPC stream (with a subprocess test that proves it).
## Quick Start
> [!IMPORTANT]
> PyMOL open-source is a **conda** package, and the server must run in a Python that can `import pymol2`.
> Install into that interpreter — do **not** use `uvx`/`fastmcp install` (they build isolated envs without PyMOL).
```bash
# 1. Create the environment (or reuse one that already has pymol-open-source)
conda env create -f env.yml # env named `pymol-mcp`
conda activate pymol-mcp
# 2. Install this package (with the optional MD bridge + dev tools)
pip install -e ".[md,dev]"
# 3. Verify
pytest -q
```
It's a standard **MCP server over stdio**, so it works with any MCP-capable client (Claude Code / Desktop,
Codex CLI, Gemini CLI, Cline, Continue, …). Point the command at the **absolute** conda interpreter so it
can `import pymol2`.
Most clients use an `mcpServers` block (Claude Code / Desktop, Gemini CLI, Cline, Continue, …):
```json
{
"mcpServers": {
"pymol": {
"command": "/absolute/path/to/conda/envs/pymol-mcp/bin/python",
"args": ["-m", "pymol_mcp"]
}
}
}
```
Codex CLI — ~/.codex/config.toml
```toml
[mcp_servers.pymol]
command = "/absolute/path/to/conda/envs/pymol-mcp/bin/python"
args = ["-m", "pymol_mcp"]
```
Prefer not to hardcode a path? Use `"command": "conda", "args": ["run", "-n", "pymol-mcp", "python", "-m", "pymol_mcp"]` instead (requires `conda` on the client's PATH). See [llms-install.md](./llms-install.md) for a full from-scratch setup.
To enable the opt-in scripting tools, add `"env": {"PYMOL_MCP_ALLOW_CODE_EXEC": "1"}` to the server entry.
Then ask your agent things like:
```
Load ./hydrate.gro, color water by F4 order parameter, and render it.
Load md.gro + traj.xtc, show CO2 guests as spheres, render frame 50.
What's the mean H-bond coordination of the water in this structure?
Load tests/fixtures/hydrate_sII.gro, identify and mark cages, hide the waters, set orthographic projection, and render.
```
## Tool Catalog
| Group | Tools |
|-------|-------|
| **Session / IO** | `load_structure` · `fetch_pdb` · `list_objects` · `get_object_info` · `reset_session` |
| **Selection** | `select` · `get_selection_info` |
| **Representation** | `show` · `hide` · `color` · `spectrum` · `set_background` |
| **View / Render** | `orient` · `zoom` · `turn` · `set_projection` · `render_image` → 🖼️ inline PNG |
| **Measurement** | `measure_distance` · `measure_angle` · `measure_dihedral` · `align` · `save_file` |
| **Trajectory / MD** | `load_trajectory` (GROMACS/DCD) · `load_trajectory_mda` (LAMMPS/NetCDF via MDAnalysis) |
| **Clathrate domain** | `identify_cages` (TRACE) · `cage_occupancy` · `mark_cages` · `hbond_network` · `order_parameter` (F3 / F4) · `chill_plus` · `mark_chill_plus` |
| **Scripting (opt-in)** | `run_pml` · `run_python` |
### PyMOLWiki Ray styles
`render_image` accepts a fixed `ray_style` profile. If you omit it, the tool uses `pymolwiki_outline`.
Use `ray_style="default"` explicitly to preserve the current PyMOL settings.
| Profile | PyMOL setting | Use |
|---|---:|---|
| `default` | unchanged | Preserve the current scene style |
| `pymolwiki_outline` | `ray_trace_mode=1` | Color with black outlines |
| `pymolwiki_bw` | `ray_trace_mode=2` | Black-and-white outline mode |
| `pymolwiki_color` | `ray_trace_mode=3` | Quantized color with black outlines |
The PyMOLWiki profiles set antialiasing to 2. They use the built-in PyMOL ray tracer.
They restore the previous ray settings after each render. Set a white background for the closest match to the PyMOLWiki examples.
`set_projection` switches the camera between **orthographic** (parallel cage edges, typical publication lattice) and **perspective**. PyMOL's default field of view is 20°; raise it (e.g. 40) to make depth obvious. `save_file` writes a PNG/PDB/PSE to disk and creates missing parent directories. `list_objects` includes CGO overlays such as `cages` and `chill_plus`.
```json
{
"selection": "all",
"width": 1600,
"height": 1200,
"ray_style": "pymolwiki_color"
}
```
## Domain: clathrate-hydrate science
Ported from a validated Rust reference implementation and re-checked against ground truth. All analysis runs in
**nanometres** with a correct fractional-coordinate **minimum-image convention** (orthorhombic *and* triclinic),
a signed `atan2` dihedral for F4, and a periodic-image KDTree for neighbour search.
- **`identify_cages`** — full TRACE cage perception: ring finding → geometric validation → constraint-propagation assembly → Euler (SEC) validation → face-count typing (5¹², 5¹²6², 5¹²6⁴, …) and an sI/sII/sH structure call.
- **`cage_occupancy`** — assign guest molecules (CO₂/CH₄) to cages and report per-type occupancy (θ_S, θ_L). Guests are detected only from residue names containing `CO2`, `CH4`, or `METH` (carbon atom). The bundled `tests/fixtures/hydrate_sI.gro` is TIP4P water only, so occupancy is zero there.
- **`mark_cages`** — draw each cage as a **wireframe polyhedron** (CGO object `cages`): cylinders on O–O edges and spheres at oxygen vertices, colored by type (5¹² cyan, 5¹²6² violet, 5¹²6⁴ red, 5¹²6⁸ blue, 4³5⁶6³ lime). Hide the water object, then `render_image` or `save_file`.
- **`mark_cages` geometry defaults** — use a bold 0.18 Å cage edge radius and a 0.38 Å vertex radius by default. Pass `edge_radius` or `vertex_radius` to override them. `cage_types` draws a subset (e.g. `["5^12 6^8"]`). Cages that wrap the periodic box can appear as fragments; that is a drawing limit, not a miscount.
- **`order_parameter`** — F4 (torsional) and F3 (three-body angular). F4 ≈ 0.7–0.95 → hydrate, ≈ 0 → liquid, ≈ −0.4 → ice Ih.
- **`hbond_network`** — water H-bond graph (O–O ≤ 0.36 nm and a donor H–O···O angle < 35°) with coordination stats.
- **`chill_plus`** — classify each water as liquid, ice, or hydrate with the CHILL+ q3 bond correlation method. It returns six class counts, per-water classes, fractions, and the unique O–O network. The default cutoff is 0.35 nm. The `state` argument uses one-based PyMOL states and defaults to 1.
- **`mark_chill_plus`** — draw the non-liquid CHILL+ O–O network as a color-coded CGO overlay named `chill_plus` for `render_image`. It uses the same one-based `state` argument.
- **Performance** — `chill_plus` uses a sparse cell list for orthorhombic cells, including 90° angle roundoff within a scale-aware machine-precision tolerance. It keeps the canonical MIC distance check. It uses a bounded-temporary-memory O(n²) fallback for real tilts. Adjacency and edge outputs still scale with the edge count. Large triclinic systems need a separate benchmark.
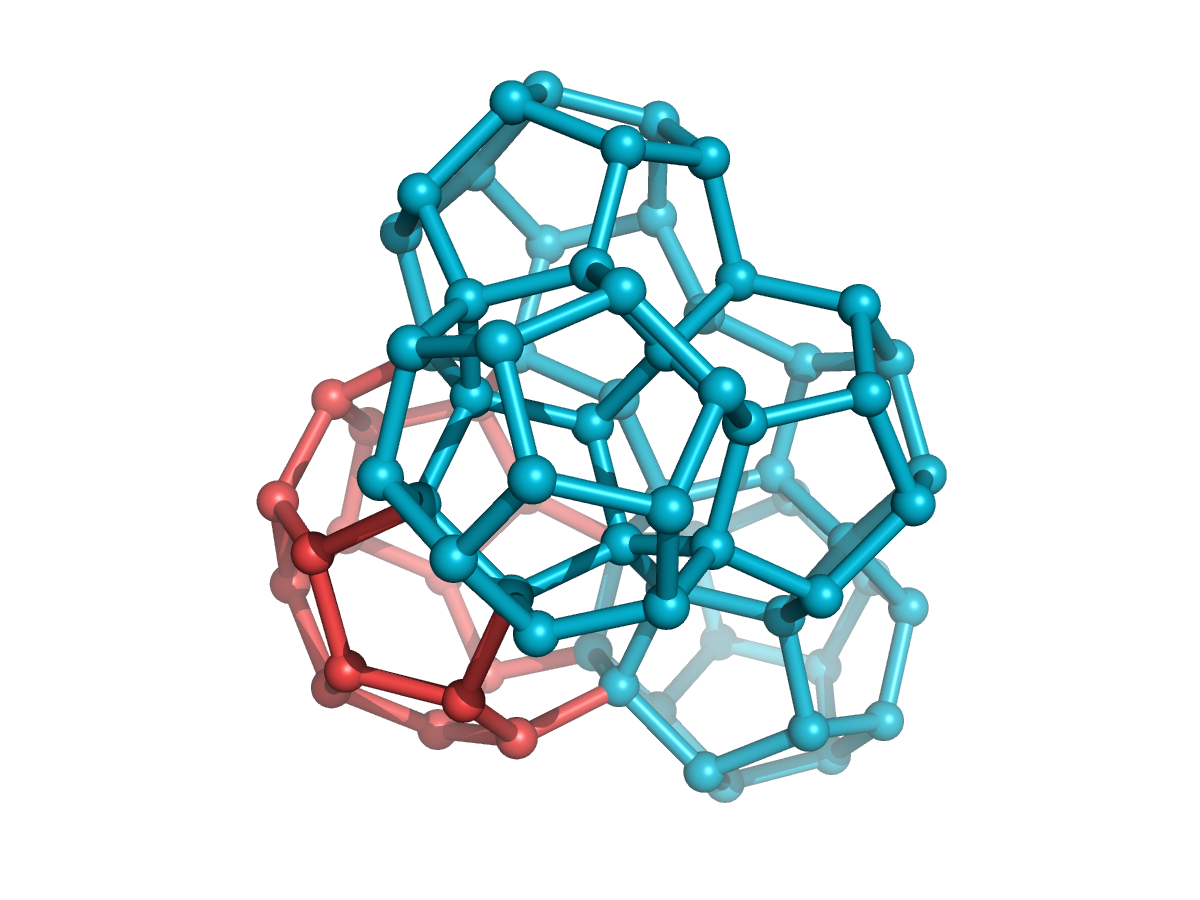
Detected sII cages drawn as wireframe polyhedra: 5¹² dodecahedra (cyan) around a 5¹²6⁴ cage (red), sharing faces.
> [!TIP]
> **Validated against ground truth:** on a structure II reference, `identify_cages` finds exactly **128 × 5¹²
> + 64 × 5¹²6⁴** cages (the textbook 2:1 sII lattice), on structure I exactly **16 × 5¹² + 48 × 5¹²6²**,
> and on structure H **48 × 5¹² + 32 × 4³5⁶6³ + 16 × 5¹²6⁸** (16 unit cells);
> F4 over the first ten waters reproduces the reference value **0.926698** exactly, F3 = 0.0028
> (hydrate-like ≤ 0.04), and the H-bond network is a perfect tetrahedral (mean coordination 4.00) framework.
A typical cage figure, exercised through the MCP tools:
```
load_structure tests/fixtures/hydrate_sII.gro (object sII)
identify_cages → structure_type sII
mark_cages → CGO object "cages"
hide water representation
set_projection orthographic (or perspective, field_of_view=40)
set_background white
render_image / save_file path.png
reset_session before the next structure
```
## How it works
```
MCP client (Claude · Codex · Gemini …)
│ stdio JSON-RPC
▼
┌───────────────────────────────────────────────┐
│ pymol-mcp (FastMCP, conda env with pymol2) │
│ • permanent stdout redirect (protocol-safe) │
│ • ONE worker thread owns + drives pymol2 │
│ • typed @mcp.tool functions │
└───────────────────────────────────────────────┘
│ cmd.* (headless) │ numpy / scipy (nm)
▼ ▼
PyMOL 3.x ── ray → PNG analysis/ (hbond, F3/F4)
coords via iterate_state
```
## Requirements
| Dependency | Required | Purpose |
|-----------|----------|---------|
| Python 3.11+ (conda) | Yes | Runtime that can `import pymol2` |
| `pymol-open-source` 3.x | Yes | The visualization engine (conda) |
| `fastmcp` 3.x, `numpy`, `scipy` | Yes | MCP server + analysis |
| `MDAnalysis` | No (extra `md`) | LAMMPS / NetCDF / xtc bridge (GPL-2.0+) |
| `ffmpeg` | No | Movie export (future) |
## Contributing
Issues and PRs welcome — see [CONTRIBUTING.md](./CONTRIBUTING.md).
## License
[MIT](./LICENSE). The optional `md` extra pulls **MDAnalysis** (GPL-2.0-or-later), imported lazily; the core
package stays MIT.